India as a Strategic Source of Global Clinical Evidence

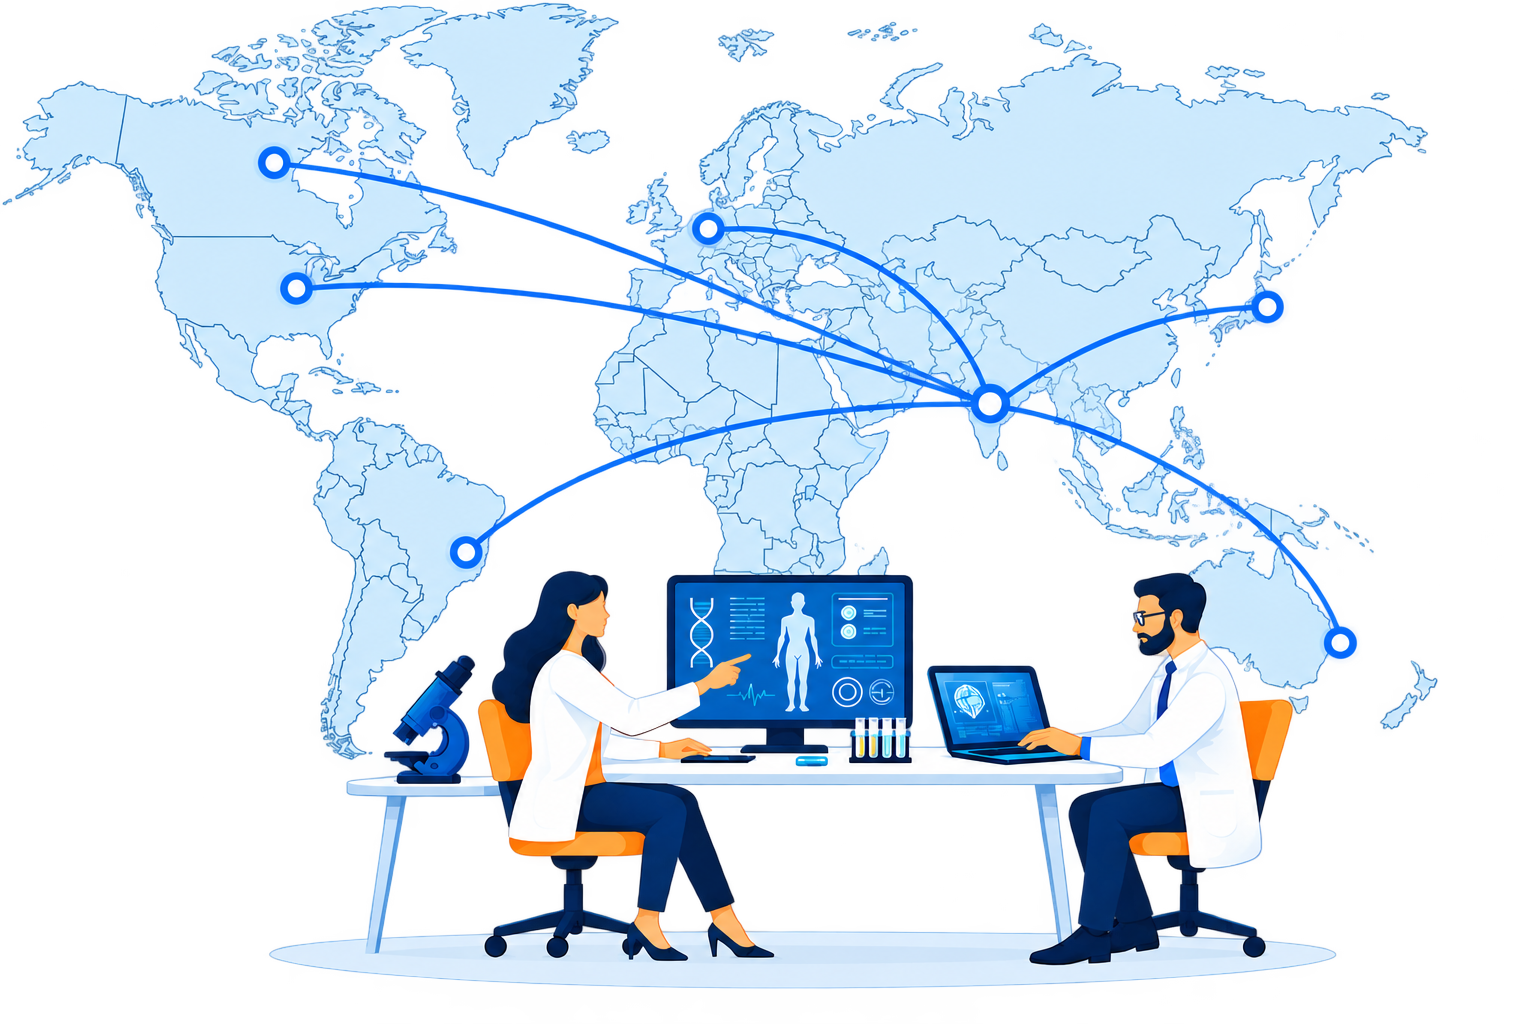
India offers access to diverse patient populations, experienced investigators, high-volume healthcare institutions, and internationally aligned clinical research infrastructure.

CliniExperts designs India-based studies with global regulatory acceptability in mind.

Global India
Us

United States

FDA

India Studies Designed to Meet FDA Expectations
ISO 14155:2020 compliant clinical investigations
ICH E6(R2) Good Clinical Practice compliance
ICH E5 ethnic factors considerations
ICH E17 multi-regional clinical trial alignment
Awareness of 21 CFR Part 812 requirements
Support for 510(k), De Novo and PMA evidence programmes
US Landmark
Europe

European Union

EU MDR & IVDR

India Evidence Supporting EU Regulatory Programmes
EU MDR Article 61 clinical evidence support
Annex XIV PMCF studies
EU IVDR PMPF programmes
ISO 14155 compliant investigations
Clinical Evaluation Report support
EUDAMED-ready documentation
Europe Landmark
Uk

United Kingdom

MHRA

India Evidence Supporting UK Market Access
UK MDR aware study planning
MHRA-aligned evidence generation
UKCA evidence support
Post-market evidence programmes
International GCP compliant investigations
UK Landmark
Japan

Japan

PMDA

India Studies Designed for PMDA Alignments
Medical Device GCP-aligned clinical investigations
Safety & performance evidence generation
Clinical endpoints & statistical strategy
Ethnic and population considerations
GCP documentation & inspection readiness
Real-world & post-market clinical evidence
Japan Landmark
Canada

Canada

Health Canada

India Evidence Supporting Medical Device Licence Applications
Class III and Class IV device evidence programmes
Health Canada clinical evidence alignment
ISO 14155 compliance
ICH GCP compliant investigations
Structured reporting for Canadian submissions
Canada Landmark
Australia

Australia

TGA

India Studies Designed for TGA

ISO 14155-compliant clinical investigations
Essential Principles-aligned evidence generation
Clinical safety & performance studies
Feasibility & pilot studies Clinical evidence gap assessment
CER & regulatory evidence support
Post-market & real-world clinical evidence
Australia Landmark

Why Global Sponsors Choose India

India
w_Global 1

Diverse Patient Populations

Accelerated recruitment and broad demographic representation
w_Global 2

International Quality Standards

Studies conducted under ISO 14155, ICH-GCP, and applicable regulatory requirements
w_Global 3

Cost-Efficient Clinical Development

High-quality evidence generation with operational efficiency.
w_Global 4

End-to-End Clinical Investigation Support

A scientifically integrated, end-to-end process from Regulatory and Ethical Approvals, Study Site operations, Data Management, Biostatistics, Medical Writing, and Safety Management, culminating in robust evidence generation and final study reporting for regulatory submission through a single strategic partner.

Frequently Asked Questions

Which class of Medical Devices require clinical investigation in India?

Medical device clinical investigation is required in India to examine all Medical Devices / in vitro diagnostic kits class A, B, C, and D; if the medical device falls within the scope of an investigational medical instrument that doesn't have a predicate, is made in India or if its a new Medical Device / in vitro diagnostic kits.

Each individual, sponsor, clinical researcher, organization, and any other investigator participating in a clinical inquiry or his agent who has permission to do so shall keep the data, record, registers, etc., for seven years from the end of the clinical investigation. The Central Licensing Authority and any other authorized officers may require such information.

Clinical investigation of medical products is the process of obtaining data to evaluate the safety and performance of medical products in their intended use. This medical device performance study includes any side effects or risks the product may cause during its use.

Form MD 22 is an application to get the grant of permission to conduct clinical investigation of medical devices without predicate devices or new ones.

A “clinical investigation plan” is a document that describes the reasons, objectives, design methodology, monitoring, and statistical considerations. It also includes the proposed analysis, monitoring, conduct, and record-keeping for the clinical investigation.

Pilot studies for medical device clinical studies or drug development are typically performed as a priority at the beginning of the process. They can be conducted when little or no clinical experience is available with the device or drug. Pilot studies refer to medical devices or drug clinical trials that gather specific essential information about a device or drug before starting the pivotal trial. These trials will be used to make precise claims about safety and efficacy. A pivotal study for a medical device is a clinical test that attempts to show the efficacy of a new medical device, and it can be used to gain marketing approval from the regulatory authority. A medical device pivotal clinical study aims to demonstrate that a novel experimental medical device is more efficient than an existing standard-of-care medical device.

The Central Licensing Authority can, if satisfied with the application, grant permission to conduct a medical device clinical investigation in Form MD 23.

Any medical device or new in vitro diagnostic medical device imported or manufactured, for the purpose of clinical investigation or clinical performance evaluation, test, evaluation, demonstration and training, shall be kept in containers bearing labels, indicating the name of the product or code number, batch or lot number, serial number wherever applicable, date of manufacture, use before date, storage conditions, name and address of the manufacturer, and the purpose for which it has been manufactured.

The license pertaining to the import for test, evaluation, clinical investigations is granted using form MD-17 by the Central Licensing Authority.

The results of medical device or in-vitro diagnostic kit clinical investigation may not be required to be submitted where the investigational medical device is approved by the regulatory authorities of either the United Kingdom or the United States of America or Australia or Canada or Japan or EU and the said device has been marketed for at least two years in that country and the Central Licencing Authority is satisfied with the data of safety, performance and pharmacovigilance of the device.


*All the above fields are mandatory
contact us 2
Enquire now